Delivery: Protein-delivered base editor preserves...
CMN Briefs
Sep. 11, 2026
“The future of gene editing will be advanced by generating clear, reliable evidence that helps developers identify and evaluate potential off-target sites with confidence”Gavin Kurgan, IDT
Every therapeutic editing programme has to answer the same regulatory question: where else in the genome does the guide RNA send the editor, and does it matter? The US Food and Drug Administration's draft guidance recommends that developers use multiple orthogonal off-target nomination methods, but offers no view on which ones, and the field has no agreed gold standard against which any of them can be scored. The result is that two assays run on the same guide can return lists that differ by an order of magnitude, with no way of knowing which sites were missed. Kyle Kinney and colleagues at Integrated DNA Technologies and the University of California, San Francisco, set out to close that gap, and they recently published their results in Nature Communications.
»The future of gene editing will be advanced by generating clear, reliable evidence that helps developers identify and evaluate potential off-target sites with confidence,« says Gavin Kurgan in a company press release. He is senior manager of bioinformatics applications development at IDT and the study's corresponding author.
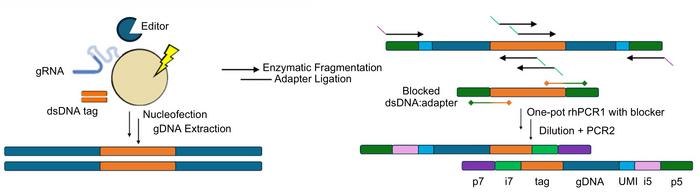
The method, UNCOVERseq, is a cell-based assay built on the familiar principle of capturing a double-stranded DNA tag at nuclease-induced breaks, but rebuilt end to end (see Figure 1). A blocking oligonucleotide removes an artefact that had been consuming 37–67 per cent of sequencing reads in the original GUIDE-seq protocol, and a new alignment and statistical pipeline replaces string matching with a glocal Needleman-Wunsch implementation. Across 48 guides, the reworked workflow nominated a median of 132 off-target sites per guide compared with 23 for GUIDE-seq – between 60 and 4,150 per cent more.
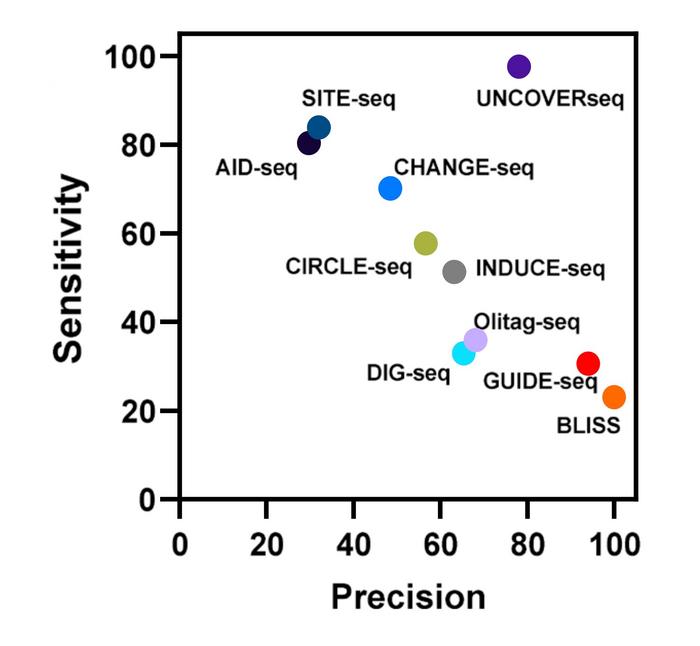
Volume alone proves nothing, so the authors built a benchmarking dataset in which nominated sites were confirmed by ultra-deep amplicon sequencing. Against that dataset, the assay recovered 97.6 per cent of confirmed off-target edits at 78 per cent precision.
The comparison is unflattering for the methods most often cited in regulatory filings (see Figure 2). GUIDE-seq recovered 35.9 per cent of true positives, BLISS 23.1 per cent and DIG-seq 33.0 per cent, while the biochemical in vitro assays SITE-seq and AID-seq reached 83.8 and 80.4 per cent sensitivity but at precision of only 32.1 and 29.7 per cent respectively. In other words, the cell-based assays in common use missed the majority of confirmable events, and the biochemical assays that found them buried the findings in long, largely false-positive lists.
The practical requirements are unusually well specified, which matters for anyone writing an assay into a development plan. UNCOVERseq needs roughly 150,000 genome equivalents of input DNA, and sequencing depth turns out to be the binding constraint: 50,000 reads per sample suffice for high-frequency events, but detecting off-targets at 0.01–0.02 per cent indel frequency requires at least two million. The authors recommend more than 500,000 reads as a minimum. Replication is not optional either – without biological triplicates, 25–40 per cent of the sites the tiering system flagged as high priority were missed or misranked.
To ask whether any of this survives the move into clinically relevant cells, the team screened 192 guides, selected six spanning the full specificity range, and edited CD34+ haematopoietic stem and progenitor cells from a healthy donor using mRNA delivery of high-fidelity S. pyogenes Cas9, adenine and cytosine base editors, and a prime editor. On-target indel editing in the stem cells ranged from 4.4 to 88.4 per cent; intended base conversion reached 16.5–75.9 per cent for the adenine base editor and 0.78–32.9 per cent for the cytosine base editor. Prime editing produced almost no on-target activity without further pegRNA optimisation and was excluded from the off-target analysis.
Between 2.4 and 34.5 per cent of nominated sites converted to confirmed editing in the stem cells, with frequencies from 0.06 to 88 per cent. The conversion rate fell as guide specificity rose, and for the highest-specificity guides the only site confirmed was the on-target – the off-target burden of a well-chosen guide in a primary cell simply drops below what current sequencing can measure. That is a reassuring finding, but it is also a measurement limit rather than a biological zero.
The most clinically pointed result concerns editor choice. Sites nominated on the basis of double-strand breaks predicted where base editors would act, with rank-order correlations of 0.77 for the adenine and 0.78 for the cytosine editor, so a single nuclease-based nomination can serve both modalities.
“We hope this work provides scientists, developers and regulators with a practical framework for evaluating off-target nomination strategies with greater confidence”Gavin Kurgan, IDT
But when the authors summed the full off-target burden per on-target event – indels, single-strand-break edits and translocations together – the high-fidelity nuclease came out ahead of the base editors. Translocations, meanwhile, were detectable for only one of the six guides, at 0.3-1.7 per cent, suggesting they are rare in healthy stem cells across guides of higher specificity.
Given that the FDA is currently weighing draft guidance on sequencing strategies and reporting for off-target editing in support of IND and licence applications, Kurgan sees the timing of UNCOVERseq as significant.
»By working across disciplines to measure sensitivity and precision against empirically confirmed editing – and by establishing the operating parameters that influence those measurements – we hope this work provides scientists, developers and regulators with a practical framework for evaluating off-target nomination strategies with greater confidence,« he says in the press release.
Some caveats are worth holding on to. Much of the nomination work is done in a promiscuous HEK293 line stably expressing Cas9, deliberately chosen to over-report, and the head-to-head performance figures rest on two guide RNAs. The field also still lacks the standardised, ACMG-style scoring system that would turn a ranked site list into a risk call, and structural events such as loss of heterozygosity and multi-kilobase deletions remain outside the assay's reach.
The study was led by Kyle Kinney and Gavin Kurgan at Integrated DNA Technologies, with the HSPC experiments performed by Kun Jia at the University of California, San Francisco. It was published in Nature Communications on 11 August 2026. IDT announced the results on the same day in a press release.
To get more CRISPR Medicine News delivered to your inbox, sign up to the free weekly CMN Newsletter here.
ArticleCMN HighlightsNewsOff-targetBase editorsCas9Prime editorsSafety